-110x110.png)
二类医疗器械产品范围
页面用于帮助采购商明确选型条件。是否可供应至特定市场,以最终证书核验、进口要求和合同约定为准。
证书不是一张图片,而是一组对应关系
采购时应把证书持有人、实际制造商、产品名称、型号、预期用途、标准、有效期与数据库状态逐项对应。
文件真实性
通过发证机构或监管数据库核验编号、状态与有效期。
型号覆盖
确认拟采购型号、配件和包装组合位于有效范围内。
标签一致
核对制造商、进口商、语言、UDI或批次追溯信息。
合同固化
把认证版本、验收标准与不合格处理写入采购合同。
先确认目标国家,再筛选合规型号
提交产品、数量、目的国、渠道与证书要求,获得可执行的采购建议。
二类医疗器械出口采购核验清单 Class II Medical Devices
以下内容用于建立可比较、可追溯的采购条件,不代表任何具体型号已经取得目标市场许可。
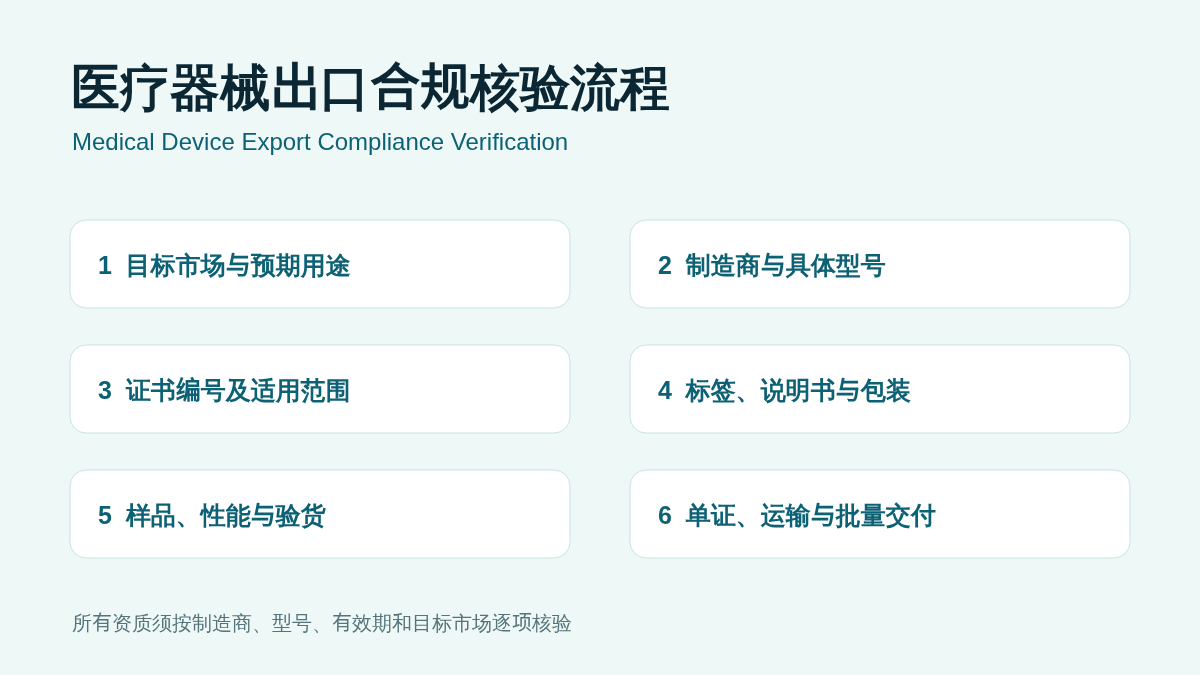
International buyer verification checklist
1. Class II Medical Devices: intended use and target market
Before quotation, define the intended use, user group, operating environment, destination country, import route, sales channel and local-language obligations for the class II medical device portfolio. These facts can affect classification, evidence, labeling and registration. A catalogue or supplier statement is not proof of market access. Verify the exact legal manufacturer, manufacturing site, brand, model, configuration and destination-market scope. Record assumptions in writing, because a change in use, user, accessory or sales country can require a new assessment even when the product appears similar.
2. Class II Medical Devices: specification baseline
Create a controlled data sheet covering intended use, device configuration, performance claims, accessories, software where applicable, labeling, regulatory route and destination-market evidence. Ask every candidate supplier to respond to the same fields and identify optional items separately. Undefined terms should be resolved before samples or quotations are compared. Final specifications must be supported by manufacturer documents, approved samples and purchase-contract requirements; this page does not confirm the specification or availability of any model.
3. Class II Medical Devices: Review performance evidence
Request current performance summaries and test reports that match the proposed model. The review should cover accuracy, precision, repeatability, interference, usability, stability, transport conditions, and lot consistency where applicable. Acceptance criteria must be agreed before sampling or inspection. Buyers should not rely only on marketing statements such as high accuracy or fast testing. Any performance claim used on packaging, websites, catalogues, or distributor materials should be supported by evidence controlled by the manufacturer and permitted in the destination market.
4. Class II Medical Devices: Verify regulatory documents
For the United States, ask for the relevant establishment, device listing, premarket submission, or exemption basis as applicable, and independently verify the exact product information through official sources. For the European market, confirm the applicable regulatory route, declaration of conformity, notified body involvement when required, certificate scope, authorized representative, UDI responsibilities, and language obligations. FDA, CE, 510(k), and other references must never be copied from a different factory, brand, family, or model. Start with the official FDA medical devices resource and obtain professional regulatory advice for the destination market.
5. Class II Medical Devices: Check labeling and packaging
Approve the product label, retail box, master carton, instructions, symbols, warnings, importer information, traceability elements, barcodes, manufacturing date, expiry date, storage conditions, and country-specific language before mass production. Private-label projects require written allocation of responsibilities for artwork approval and change control. Confirm whether the quotation includes printing plates, color boxes, manuals, control solution, lancets, batteries, and export cartons. Packaging validation should reflect the shipment method and distribution environment rather than only the appearance of the retail package.
6. Class II Medical Devices: samples, production and inspection
Use an approved reference sample and signed specification sheet for the class II medical device portfolio. The purchase contract should identify the manufacturer, model or reference, included items, packaging revision, quantity tolerance, inspection standard and required documents. During production, control critical materials and any proposed substitution. Pre-shipment inspection can cover model identity, essential functions, accessories, warnings, packaging, documentation and risk-based performance checks. Inspection helps detect differences from the approved sample but does not replace regulatory approval, qualified laboratory work or destination-market review.
7. Class II Medical Devices: logistics and after-sales support
Confirm product-specific shelf life, battery or consumable conditions, transport limits and warehouse controls. Record carton dimensions, gross weight, pallet plan, shipping marks, handling instructions and any battery or dangerous-goods considerations. For recurring orders, plan safety stock for monitoring devices, rehabilitation products, accessories, consumables, spare parts and approved packaging. Agree how damage, shortage, complaints, replacement parts, corrective action and possible recalls will be handled. Distribution and warehouse controls should preserve identification, traceability and approved storage conditions through final delivery.
8. Class II Medical Devices: supplier identity and responsibility
Identify the legal manufacturer, production address, specification owner, brand owner, exporter and any authorized representative for the class II medical device portfolio. The invoicing company may differ from the manufacturer shown on regulatory documents. In OEM or private-label projects, document who controls design, risk management, performance evidence, labeling, change control, vigilance and post-market responsibilities. Changes to the factory, brand, material, component, software or model scope require documented review rather than an assumption that earlier evidence still applies.
9. Class II Medical Devices: options and configuration control
List every selectable configuration, accessory, consumable, language, package and private-label option for the class II medical device portfolio. Identify which combinations are covered by the quoted documents and which require separate evaluation. If software, connectivity, batteries or powered accessories apply, confirm versions, interfaces, update responsibilities and compatibility. The supplier should control configuration changes and notify the buyer before substitution. A commercial option must not be represented as approved for the destination market until its exact scope has been verified.
10. Class II Medical Devices: sample evaluation protocol
Prepare a written sample protocol for the class II medical device portfolio. Record manufacturer, model, lot or serial information, configuration, accessories, document revision, packaging version and receipt date. Evaluate model identity, essential functions, accessories, warnings, packaging, documentation and risk-based performance checks. Where performance or laboratory evaluation is required, use a competent organization and documented method. Retain approved samples and evidence as production references. Sample acceptance applies only to the evaluated configuration and does not automatically approve future changes or substitutions.
11. Class II Medical Devices: quotation comparison
Require every supplier to quote the same specification and separate the costs of monitoring devices, rehabilitation products, accessories, consumables, spare parts and approved packaging. Record minimum order quantity, lead time, payment terms, warranty, replacement policy, production capacity, sample cost, inspection, tooling, artwork, documents and freight terms. Compare total landed and lifecycle cost rather than only the unit price. Exclusions, optional items and assumptions should be written clearly so that commercial comparison does not conceal a missing accessory, short shelf life, unsuitable package or unsupported claim.
12. Class II Medical Devices: traceability and change control
Define traceability for the class II medical device portfolio from finished lot or serial number to production and supplied materials. Purchase documents should require advance notice of material, component, software, packaging, manufacturing-site and regulatory changes. Decide which changes require new documents, samples, tests, artwork or commercial approval. Maintain a controlled file of quotations, specifications, samples, certificates, declarations, reports, labels, inspection results, shipping records, complaints and change notices. Revision dates should show which evidence applied to each shipment.
13. Class II Medical Devices: complaint and recall readiness
Agree a route for complaints, technical investigation, replacement decisions, reporting, corrective action and recall involving the class II medical device portfolio. Required records may include model, lot or serial identifiers, photographs, storage history, user description, accessories and returned samples. Responsibilities and timelines depend on the destination market and contractual roles. Sales staff should not make unsupported medical conclusions. The legal manufacturer or qualified regulatory party should assess reportability, corrective action and any field safety obligations.
14. Class II Medical Devices: shipment release file
Before shipment, check the approved order, specification, quantity, production identification, expiry or service-life information where applicable, carton marks, inspection report, invoice, packing list, origin documents and destination-specific evidence for the class II medical device portfolio. Compare manufacturer and model information across labels, instructions, declarations, certificates and shipping records. Resolve inconsistencies before release. For consolidation or overseas warehousing, define storage, stock rotation, damage handling and traceability. Release should follow a completed checklist rather than an informal message.
15. Class II Medical Devices: information required for an accurate inquiry
A useful inquiry for the class II medical device portfolio states destination country, intended user, distribution channel, estimated quantity, target delivery, required configuration, language, packaging, accessories, private-label needs and documents to be reviewed. Also state whether samples, third-party testing, factory audit, pre-shipment inspection, palletization or overseas warehouse support are required. When these conditions are unknown, any response is preliminary. Final selection, pricing, lead time and suitability require confirmation with the manufacturer and buyer before a binding order.
16. Class II Medical Devices: factory capability review
Review whether the proposed manufacturing site controls incoming materials, production, environmental conditions where relevant, in-process checks, release testing, nonconforming output and record retention for the class II medical device portfolio. A company presentation is not evidence that the quoted model is made at the stated site. Identify critical subcontractors and quality agreements. Compare capacity claims with equipment, validated processes, staffing, material supply and realistic lead times. Factory review should match the exact product and service scope being purchased.
17. Class II Medical Devices: quality agreement
A quality agreement can define document ownership, acceptance criteria, lot release, retention samples, complaint cooperation, change notification, audit rights, confidentiality, and record availability. It should be consistent with the commercial contract and regulatory roles. The buyer should identify which documents are required with every shipment and which may be reviewed periodically. Where translations are used, establish the controlling language and an approval route. Do not treat a certificate as a complete quality agreement. Certificates describe a limited scope and period, while the agreement controls day-to-day responsibilities for the specific products and transactions.
18. Class II Medical Devices: shelf life, service life and stability
Confirm product-specific shelf life, battery or consumable conditions, transport limits and warehouse controls. Ask how claimed periods and limits were established, which conditions apply after opening or assembly, and what remaining validity will be delivered. Evaluate expected transport and warehouse conditions, including temperature, humidity, light, vibration and stacking as applicable. An external carton alone does not prove protection through sea, air, road or warehouse handling. Excursions, damage and expired evidence require documented assessment rather than automatic acceptance.
19. Class II Medical Devices: localization control
Localization of the class II medical device portfolio can include labels, instructions, warnings, symbols, units, date formats, customer-service details, packaging claims, quick guides, software or screen text where applicable. Use qualified technical review and maintain approved artwork with revision history. Compare production samples against the controlled files. Machine translation without review can create misleading or unsafe information. Agree who owns translation accuracy, regulatory review, printing approval and removal of obsolete material after changes.
20. Class II Medical Devices: tender and distributor documentation
Institutional tenders and distributor onboarding often require a structured document package. Prepare a list before quotation that may include company registration, manufacturing information, quality certificates, product declarations, applicable market authorizations, technical specifications, test summaries, labeling, instructions, warranty terms, after-sales procedure, and references permitted by law. Each document should show its issuer, model scope, revision, date, and validity where relevant. Avoid mixing documents from different configurations. If a tender requirement cannot be met, record the gap clearly rather than substituting an unrelated certificate or changing the wording of the requirement.
21. Class II Medical Devices: inspection sampling and acceptance
Set inspection scope for the class II medical device portfolio according to risk, order size, supplier history and agreed specifications. Define lot formation, sample size, acceptance criteria and treatment of critical, major and minor defects. Checks may include model identity, essential functions, accessories, warnings, packaging, documentation and risk-based performance checks. Any destructive, clinical or laboratory assessment requires suitable methods and competence. The report should identify the lot, sample, method, findings, photographs and disposition and must not be presented as a regulatory certificate.
22. Class II Medical Devices: payment and commercial risk
Commercial terms should protect both parties without weakening quality controls. Align deposits, production milestones, balance payment, inspection, document approval, and shipment release. Clarify the Incoterm, nominated carrier, insurance, bank charges, customs responsibility, and consequences of delay or failed inspection. If a supplier requests payment to a different company, verify the relationship and contractual authority. Avoid approving production before specifications and artwork are frozen. A clear change-order process helps prevent disputes when quantity, packaging, accessories, language, or delivery plans change after quotation.
23. Class II Medical Devices: ongoing supplier monitoring
Supplier approval for the class II medical device portfolio is not permanent. Monitor delivery, document expiry, inspection results, complaint trends, corrective action, change notices, responsiveness and lot consistency. Establish review intervals based on risk and purchase volume. New sites, materials, components, software, accessories, packaging or market authorizations can trigger reassessment. Objective records and measurable expectations support stable supply and earlier detection of problems; repeated exceptions without investigation increase commercial and regulatory risk.
24. Class II Medical Devices: boundaries of this sourcing service
The sourcing process can coordinate supplier identification, document collection, quotation comparison, sampling, inspection, packaging, logistics, and communication. It does not replace the legal manufacturer, accredited laboratory, notified body, regulator, importer, authorized representative, or professional legal advice. Final classification and market-access decisions must be made for the exact product and destination. No wording on this page should be interpreted as confirmation that a particular model is approved, registered, cleared, certified, or suitable for a specific clinical purpose until the supporting evidence has been reviewed and accepted.
25. Class II Medical Devices: final decision record
Before placing the order, create a decision record that summarizes the intended market, selected manufacturer and model, supporting documents, approved samples, open risks, responsible parties, inspection plan, delivery terms, and final approvals. Link each conclusion to a dated source document. Record unresolved assumptions and the person authorized to accept them. This final review helps prevent a commercial quotation from being mistaken for technical or regulatory approval. It also provides a useful baseline for repeat orders, supplier changes, customer questions, and post-market investigations.
26. Class II Medical Devices: controlled document index
Create a document index that lists every file received for the proposed product, including issuer, document number, revision, date, expiry where applicable, covered manufacturer, covered site, model scope, language and verification status. Link conclusions to the exact file rather than a screenshot or sales message. Mark superseded documents clearly and restrict uncontrolled copies. The index should identify missing information, responsible reviewers and the date of the next review. A complete index reduces delays during customer qualification, customs questions, tender preparation, repeat orders and post-market investigation while preventing unrelated evidence from being attached to the wrong product.
27. Class II Medical Devices: sample-to-order consistency
Compare quotation, approved sample, specification, artwork, production order, inspection record and shipping documents before release. Names, references, accessories, materials, claims and package quantities should remain consistent. If a difference is accepted, record who approved it, why it was acceptable and whether new technical or regulatory review was required. Do not allow a production convenience to become an undocumented specification change. For repeat orders, compare the new lot with the controlled baseline and previous corrective actions. This discipline supports predictable quality and gives both buyer and supplier a clear record when questions arise.
28. Class II Medical Devices: claims and marketing governance
Review every performance, clinical, safety, durability and compatibility claim before it appears in catalogues, online listings, packaging, manuals or distributor materials. The wording should match available evidence and destination-market rules. Avoid absolute claims, unsupported comparisons and statements copied from another manufacturer or model. Translations should preserve the approved meaning. Keep a claims matrix that identifies the statement, supporting evidence, approved markets, media where it may be used and responsible reviewer. Commercial teams should use only controlled material and request review before creating new claims for campaigns, tenders or customer presentations.
29. Class II Medical Devices: warehouse and stock governance
Document receiving inspection, location control, storage conditions, lot or serial traceability, first-expiry-first-out rules where applicable, damaged-stock segregation, cycle counting, picking verification and dispatch records. Warehouse staff should be able to identify product status and prevent obsolete, recalled, expired or unapproved stock from shipping. When overseas inventory is planned, define replenishment triggers, safety stock, replacement parts and responsibility for local returns. Audit records should connect each outbound shipment with its source lot and approved documentation. Warehouse capability must be verified rather than assumed from a general statement that overseas stock is available.
30. Class II Medical Devices: final inquiry checklist
Send the destination country, intended use, buyer type, annual forecast, first-order quantity, required delivery date, preferred configuration, packaging language, private-label request, inspection needs, warehouse plan and document checklist in one structured inquiry. Ask the supplier to identify exceptions and assumptions instead of answering only with a price. The resulting quotation should be version controlled and linked to the offered manufacturer and model. Before confirmation, close every critical gap or record an authorized risk decision. This approach produces a clearer comparison, a more reliable contract and a faster route from sample evaluation to compliant commercial delivery.